2026-10-13 ピッツバーグ大学
<関連情報>
- https://www.engineering.pitt.edu/news/2026/finding-the-right-starting-place-for-chemistrys-most-difficult-molecules/
- https://www.nature.com/articles/s41467-026-77540-x
Kohn-Sham密度エンコーディングを用いた強相関分子に対する結合クラスター理論の改良 Improving coupled cluster theory for strongly correlated molecules with Kohn-Sham density encoding
Abdulrahman Y. Zamani, Barbaro Zulueta, Andrew M. Ricciuti, John A. Keith & Kay Carter-Fenk
Nature Communications Published:11 September 2026
DOI:https://doi.org/10.1038/s41467-026-77540-x Early provide
Abstract
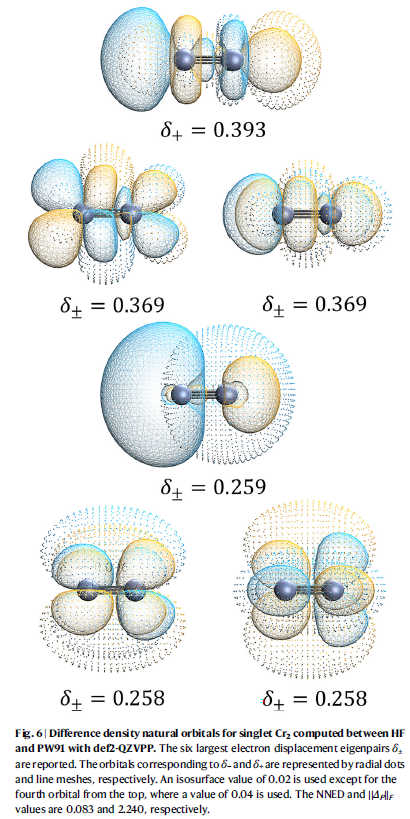
Coupled cluster theory using a Kohn-Sham density functional reference can dramatically outperform the standard Hartree-Fock-based approach for strongly correlated systems, but the origin of these improvements has remained unclear. We show these improvements arise from differences in the one-particle density as encoded into the Fock matrix, not from the nature of the orbitals themselves, as is commonly assumed. Exploiting this insight, Kohn-Sham references enable near-chemical accuracy for electronic and thermochemical properties of transition metal dimers and main group compounds. Notably, Kohn-Sham coupled cluster qualitatively recovers the entire chromium dimer potential energy surface, a notorious failure case for conventional methods. We further introduce a density difference diagnostic that detects multireference character and guides the selection of optimal references at mean-field cost. Together, these results establish a practical route to treating strong correlation within quantum chemistry’s “gold standard” framework, with immediate implications for machine learning potential development and materials research, which rely heavily on density functional theory.